


crystALL manuscript: arXiv link pending
Example Publications using crystALL:



crystALL is an open-source data-driven framework for atomic structure prediction and materials discovery, designed to address the critical gap between computational predictions and experimental observations. Unlike methods focused solely on stable ground states, crystALL systematically embraces structural diversity and compositional uncertainty, enabling the discovery, modeling, and validation of both stable and metastable crystal configurations, going beyond more common data-driven discovery approaches that prioritize only the lowest-energy structures often failing to explain experimental observations or propose structures stable under conditions of interest.
By leveraging millions of relaxed and experimental structures from the MPDD Database and OPTIMADE API, crystALL rapidly generates wide ensembles of candidate configurations for either target compositions or compositional ranges, with the latter being a unique strength of crystALL+MPDD combination. These candidates undergo all permutations of atomic substitution, advanced machine learning-based featurization, clustering, and multi-model property prediction, supporting robust selection and direct ab initio or experimental validation (in a typical use case).
This is just a quick introduction to the main features and workflow of crystALL. Please refer to the crystALL journal article for a complete, yet still concise, description of the framework, motivation behind it, and the underlying methodology.
On the high-level, crystALL allows you to take a chemical composition/s of interest, whether you measured it experimentally or hypothesized it, and then systematically explore them with minimal bias to form an approximately ergodic ensemble of candidates by feature space analysis and evaluation over any metric of interest to fulfill your research objectives. This description is not vague, but rather emphasizes the framework's ability to fit different needs. These can range from discovering crystal structure of previously unexplained experimental observations to high-throughput screening for novel materials exhibiting certain topological characteristics (ongoing work). Overall, crystALL is a great fit for both experimentalists and computational researchers aiming to accelerate discovery of novel materials in underexplored regions of chemical space or cases escaping the generative learning or conventional computational approaches focusing only on the most probable answers.
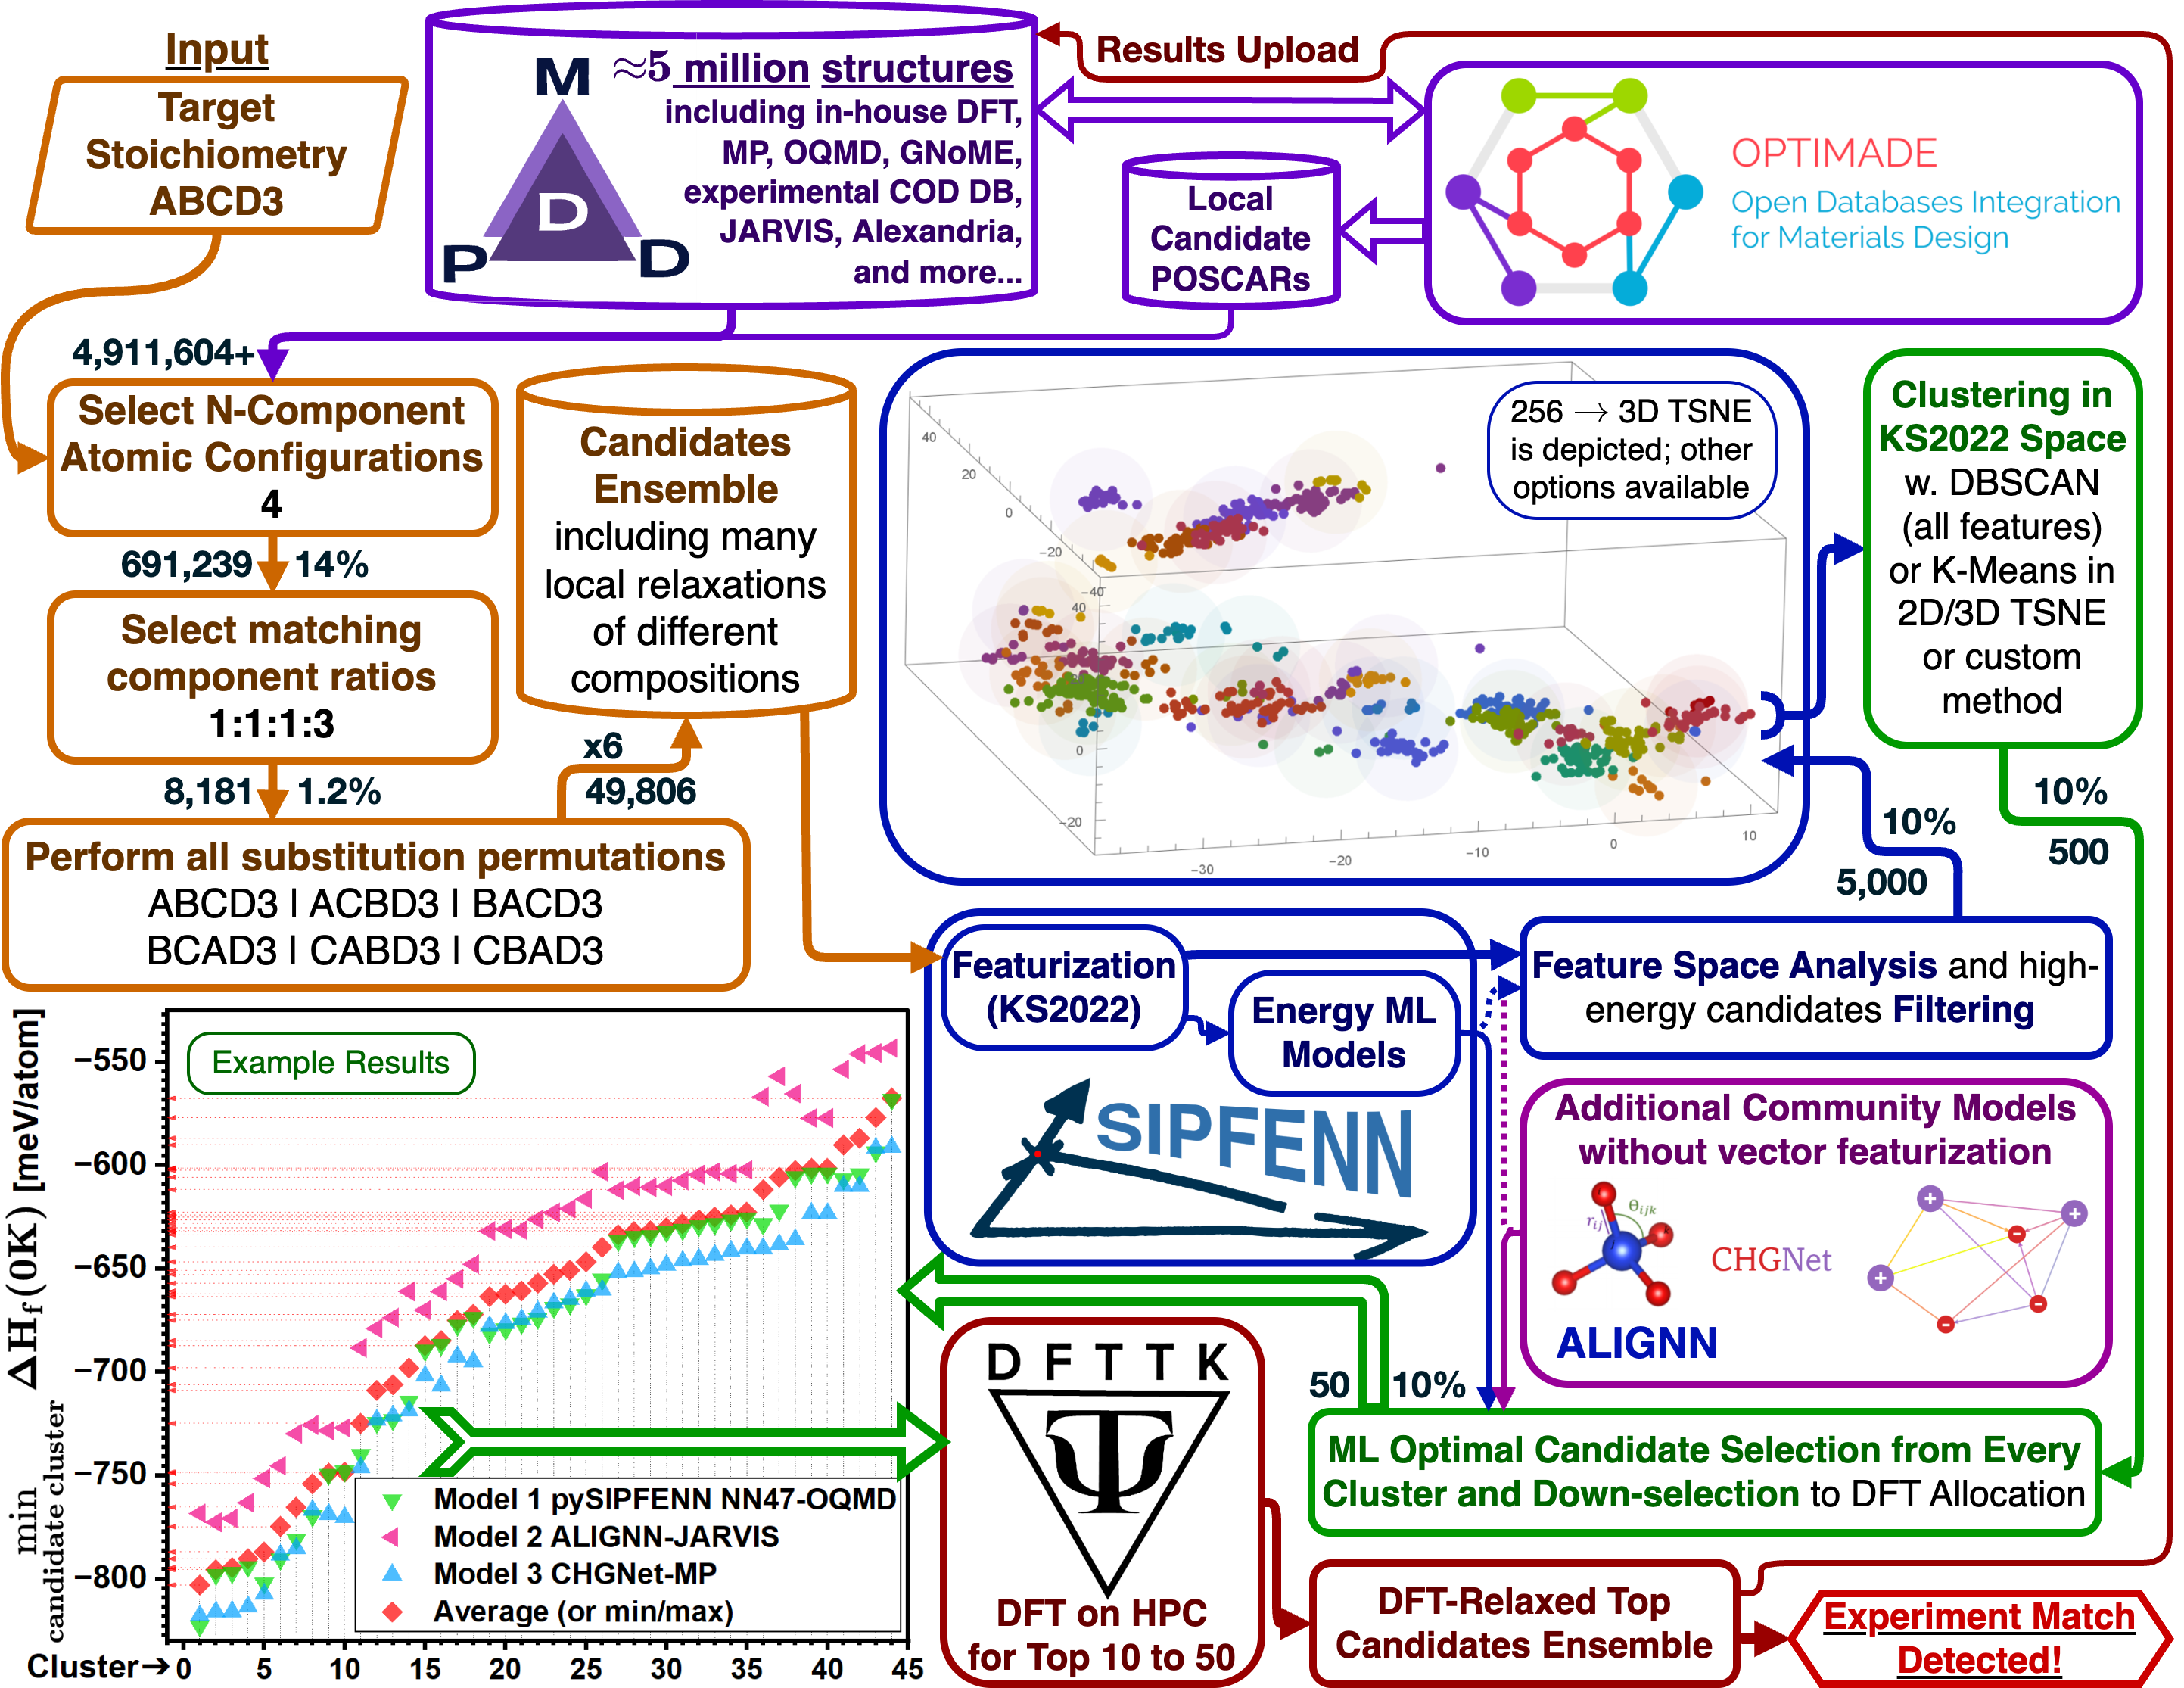
In a typical workflow, depicted in the figure below for a single target composition ("compound"), crystALL systematically generates, featurizes, and evaluates candidate crystal structures to identify the most promising configurations. It starts by gathering prototype structures (provided via MPDD / OPTIMADE API or local files) and performing all permutations of substitution into the target composition (orange). Then, it featurizes the candidates with pySIPFENN's (python toolset for Structure-Informed Property and Feature Engineering with Neural Networks) KS2022 (by default) to then analyze the embeddings (blue) while predicting properties of interest with ML models from pySIPFENN and community (purple). Filtering and clustering in the feature space are used to select top candidates from unique structures/energy basins (green), and final DFT relaxations leading to match with experimental observations (red).
The example depicts a dramatic filtering cascade - 5M structures from MPDD are harnessed to generate around 50k ABCD3 candidates, then ML-filtered to 5k in 500 clusters, culminating in DFT calculations for the top 50 cluster representatives to construct an approximately ergodic ensemble and match experimental observations; while feedback loop to MPDD creates a self-improving ecosystem for materials discovery that grows with crystALL use.
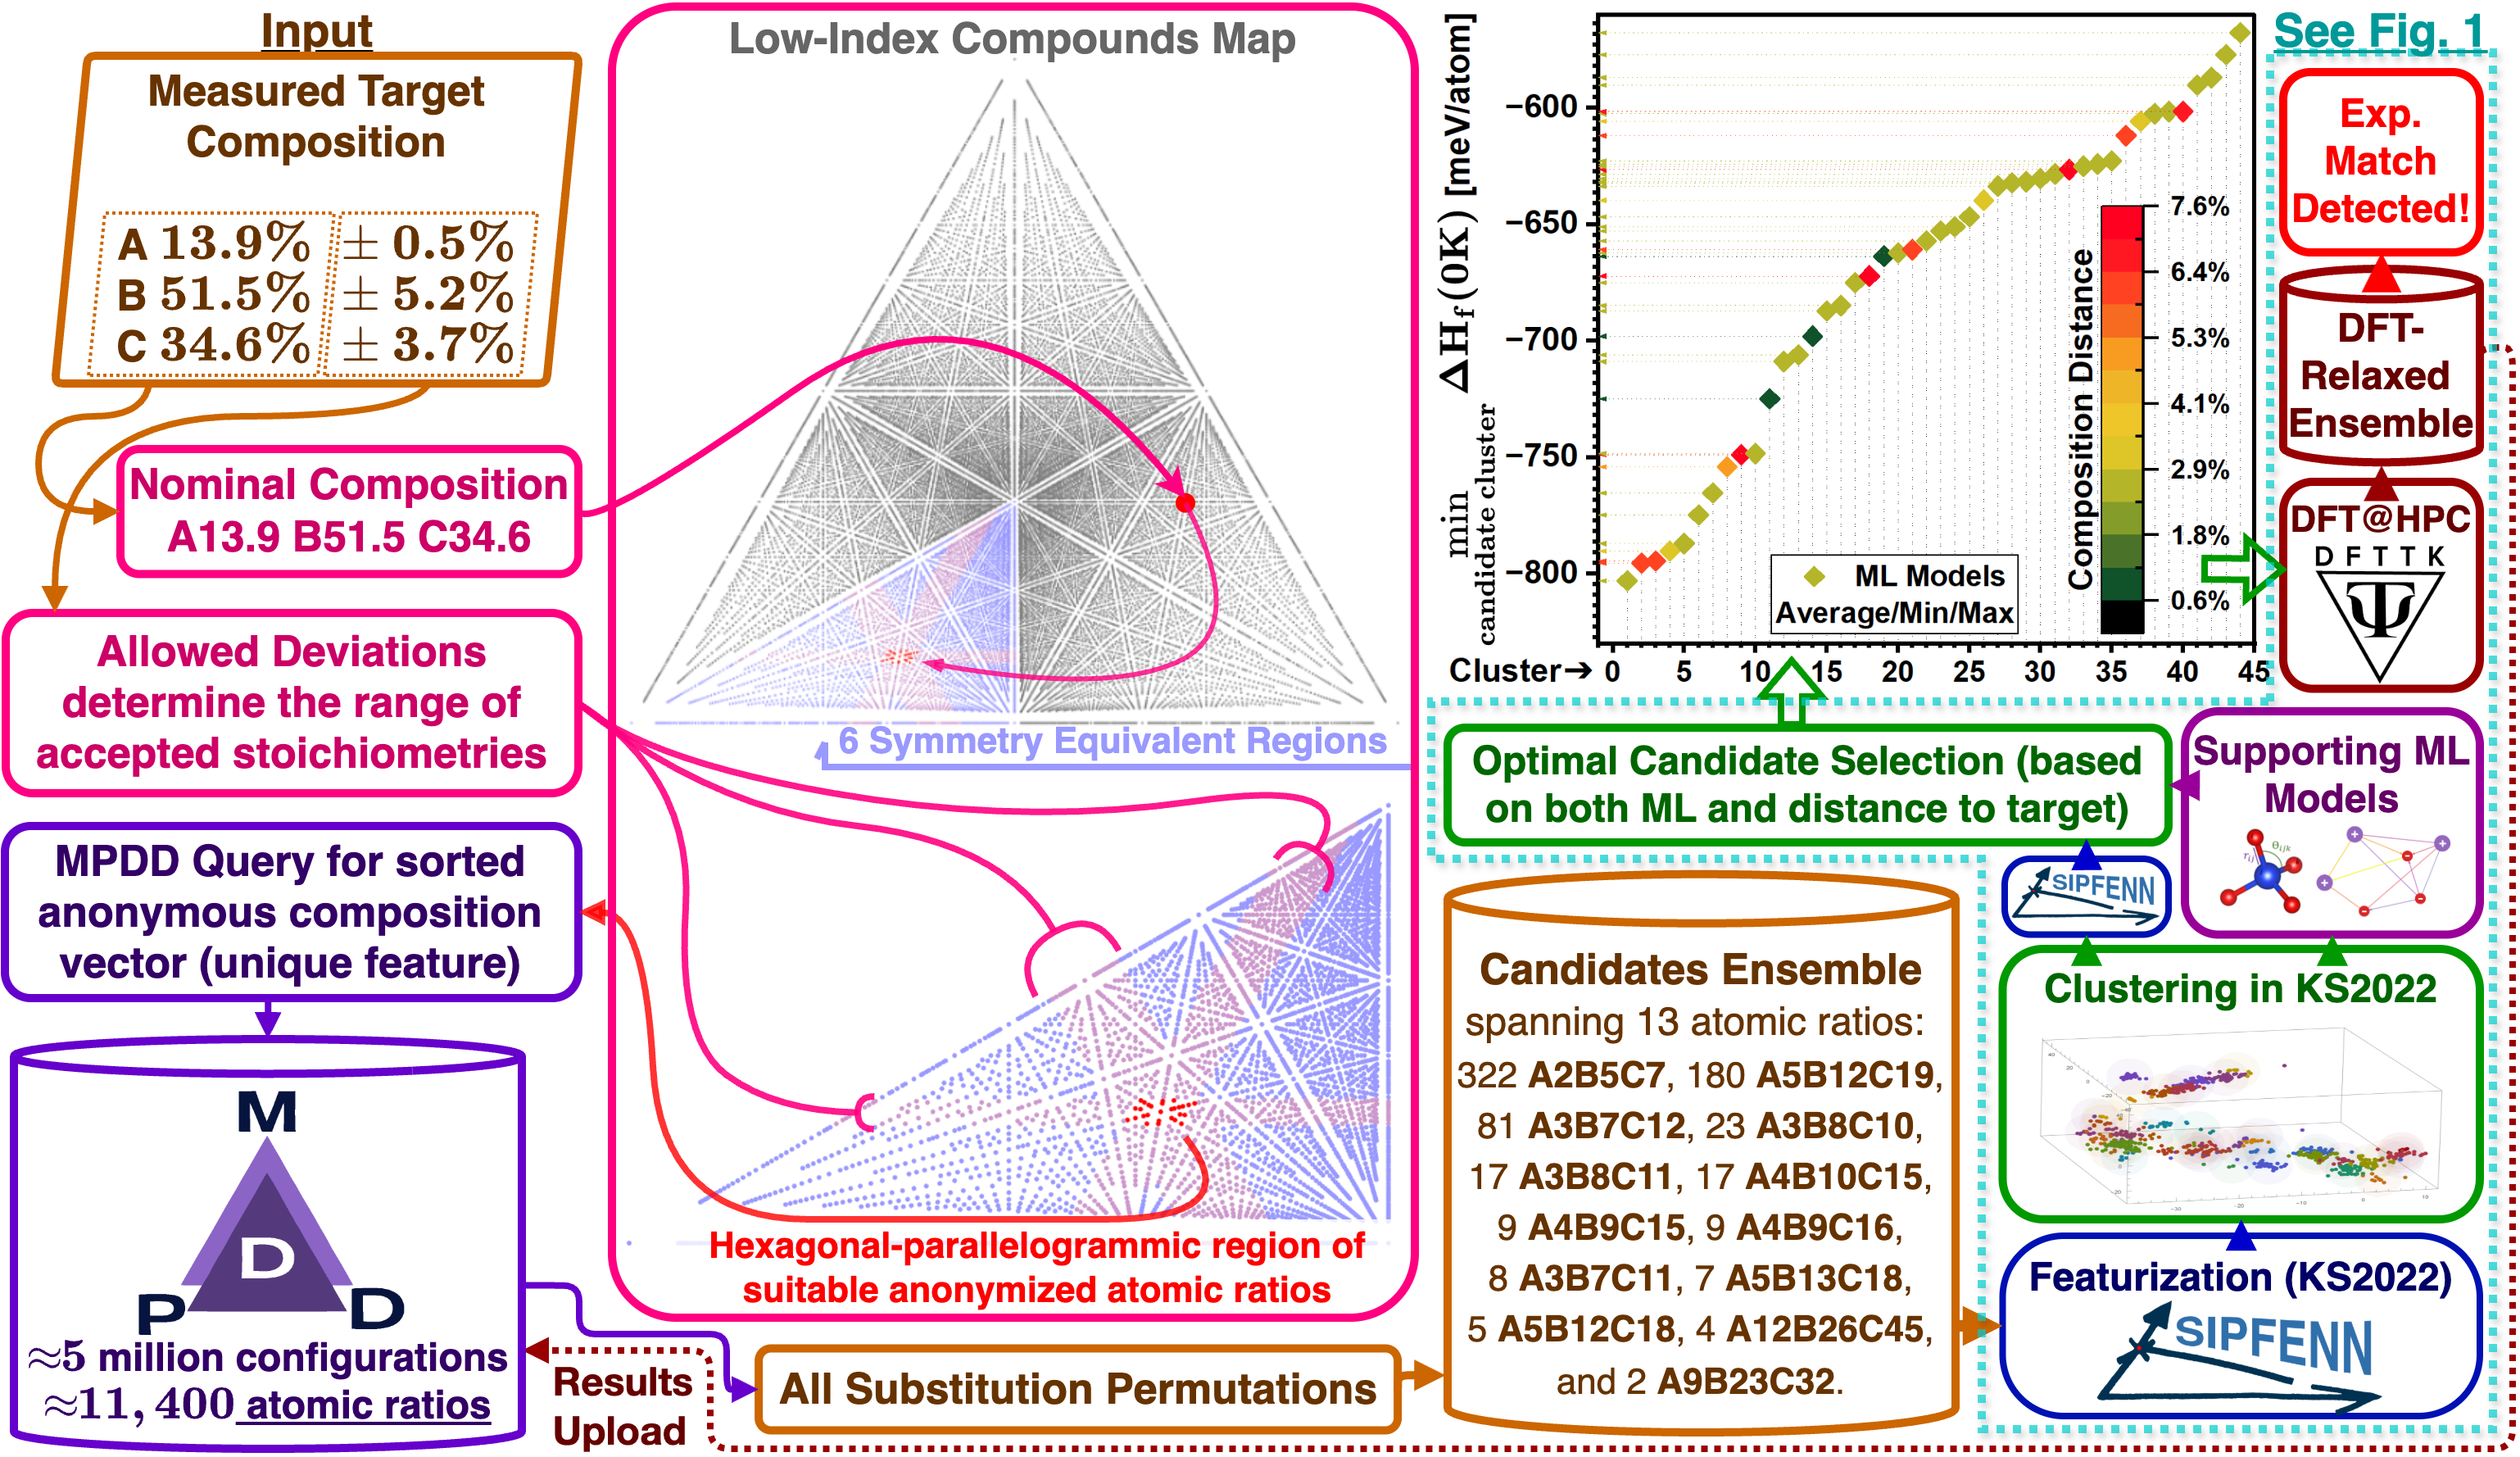
One of the unique features within crystALL, mentioned in the summary and explored in detail in the paper, is the ability to perform searches over compositional ranges, positioning it as an invaluable tool for interpreting real measurements of compositions, without assuming that molar ratios are low integers - a very common pitfall researchers fall into. As illustrated below, when combined with MPDD, crystALL accepts an uncertainty range for each element individually and systematically explores all prototypical structures within the specified compositional subspace, covering both low- and high-index anonymous formulas.
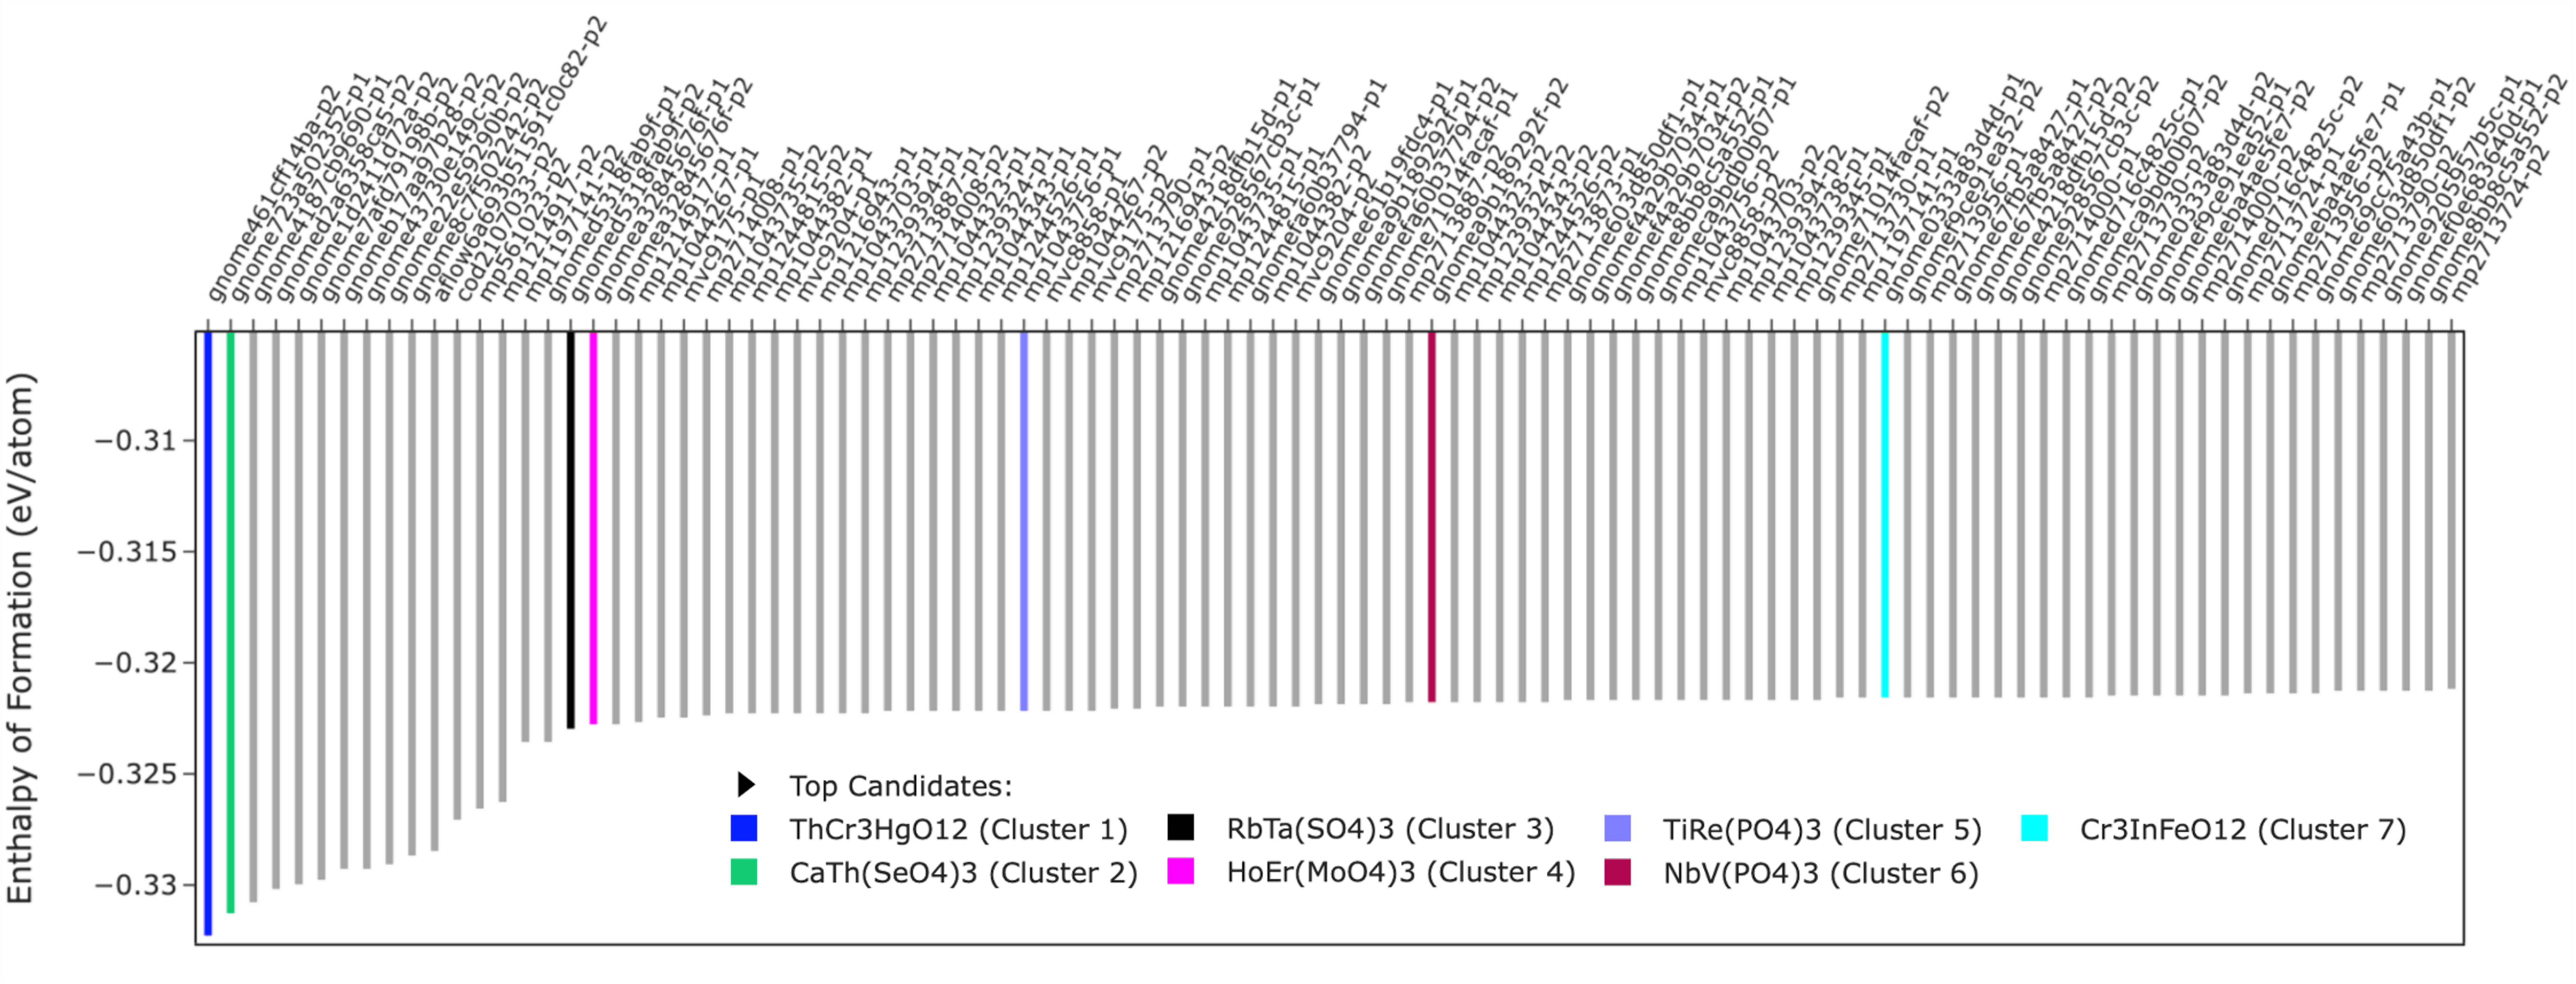
To illustrate the scoring step, the bar plot below shows formation energies sorted from lowest to highest for a focused ensemble of 100 NaMg(BH4)3 prototypes. This view helps rapidly identify low‑energy regions in the ensemble and informs which clusters deserve further attention.
Property prediction in crystALL is modular. pySIPFENN models are available natively and external calculators plug in through adapter classes. ALIGNN and CHGNet are provided as examples of such adapters: MPDD-ALIGNN exposes multi‑head formation energy outputs (based on MP, DFT-JARVIS, and Alexandria) as well as 7 other properties run by default (see MPDD-ALIGNN/alignn/config.yaml or MPDD Database (mpdd.org)), which can be filtered according to user request, and CHGNet reports static and relaxed energies computed independently so users can skip relaxation when it is not needed. The adapters accept an explicit list of objectives, enabling selective computation and reduced runtime for expensive predictors.
After featurization and property assignment, crystALL scales the 256-dimensional KS2022 descriptor vectors, projects them into 2D and 3D t-SNE manifolds for visualization, and partitions the prototype ensemble into structural basins—either directly in the full 256D scaled KS2022 feature space via DBSCAN (avoiding low-dimensional projection distortion) or on the 2D t-SNE manifold via KMeans (with automatic silhouette-based cluster-count selection when n_clusters is omitted). The figure below shows the 2D t-SNE embedding for the same NaMg(BH4)3 set: seven clusters are recovered and correspond to distinct structural families (for example, clusters 1 and 2 are enriched in hexagonal motifs, cluster 3 in triclinic arrangements, clusters 4, 5, and 7 in trigonal lattices, and cluster 6 contains perovskite-type geometries). This separation illustrates how descriptor-based clustering disentangles fundamentally different local chemistries and coordination geometries.
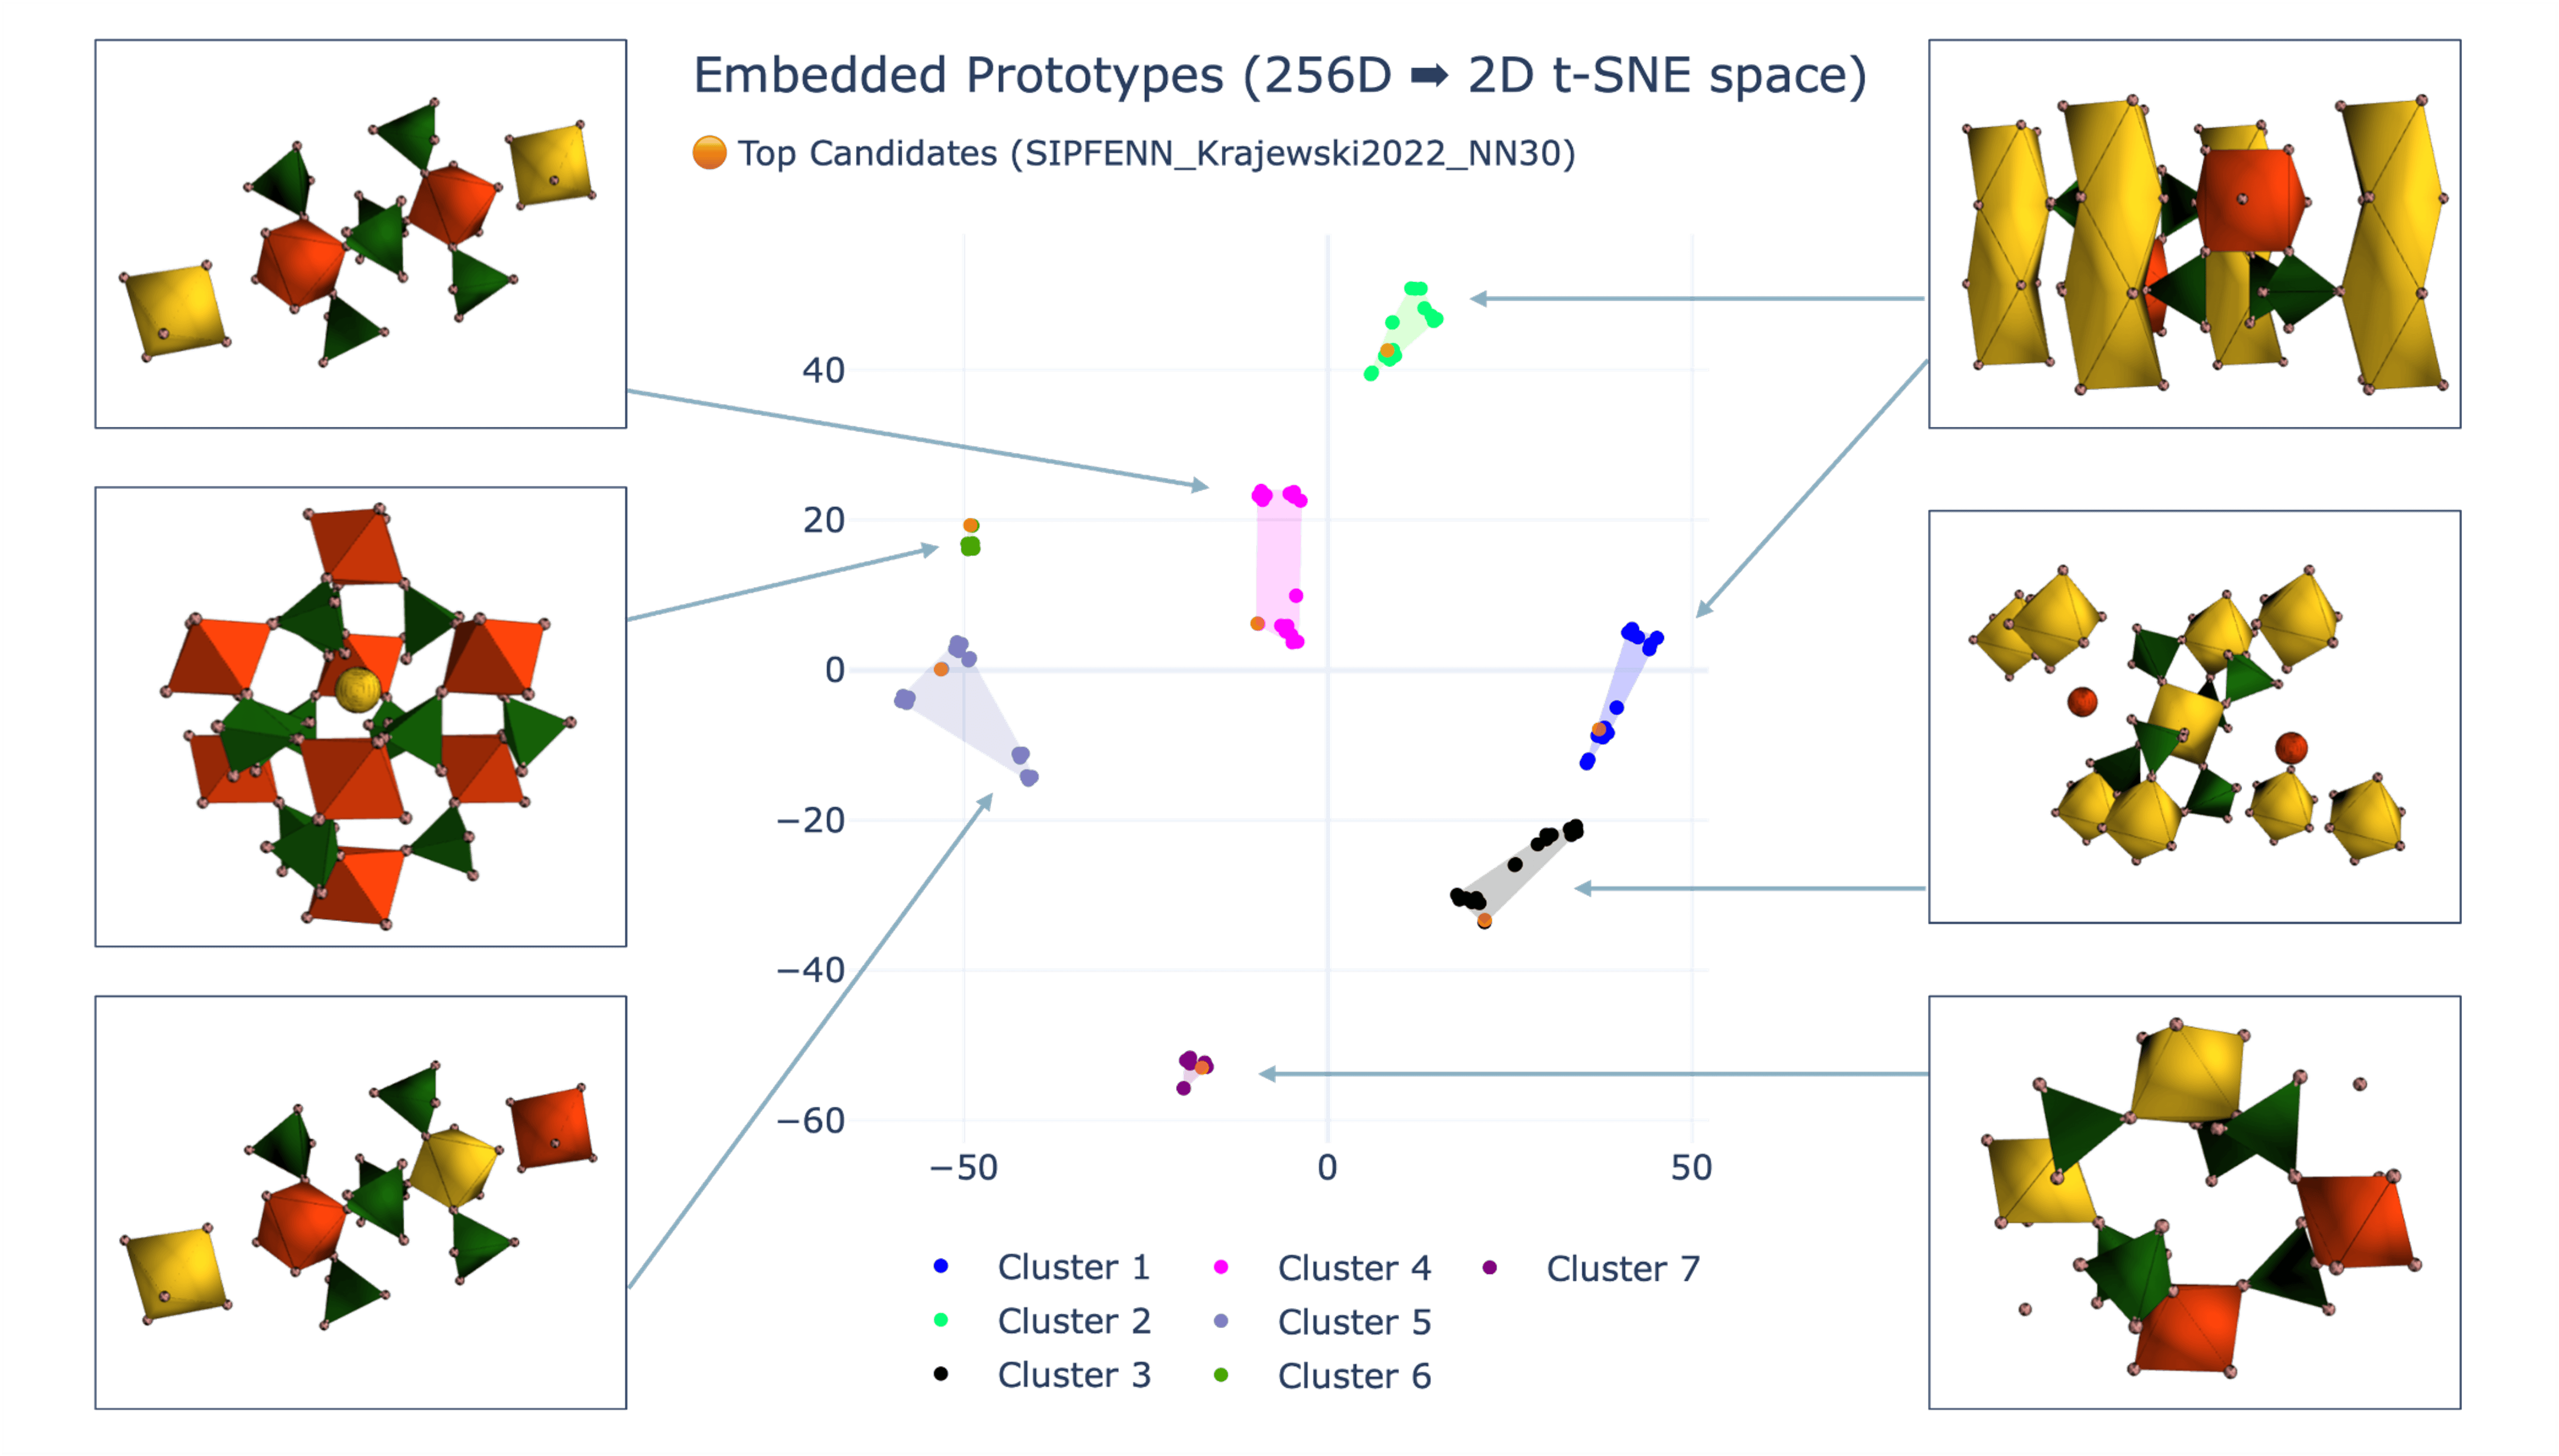
The final selection step produces Top Candidates by selecting the lowest‑energy representative from each cluster for every available objective; these candidates provide a compact, diverse set for higher‑accuracy DFT relaxation or experimental validation. Outputs are exportable as CSV (all-prototypes and top-candidates), and an optional MPDD convex‑hull check (energy‑above‑hull) can be performed to further prioritize thermodynamically plausible entries.
For detailed account of changes, please consult the GitHub changelog.
v0.5.2- Updated compatibility with the latest versions of Python and key dependencies.v0.5.1- Improved MPDD querying capabilities and documentation.v0.5.0- Major re-write of all code except for the corecrystALLlogic, turning it into a user-ready tool with additional ML models, polished architecture, user-friendly tutorials, and improved documentation.v0.4.0- Initial private GitHub release of crystALL, separated from MPDD repository; as used in 10.1016/j.actamat.2021.117448 or 10.1038/s41598-021-03578-0.
Installing crystALL on your machine can be done via the PyPI package repository, or by cloning from GitHub directly, following the instructions below. You can also effortlessly run it on the cloud by clicking here to launch a pre-configured GitHub Codespaces environment with tutorials ready to go.
While not required, it is recommended to first set up a virtual environment using micromamba, miniconda, or venv. This ensures that (a) one of the required versions of Python (3.11+) is used and (b) there are no dependency conflicts. If you have Conda installed on your system (see micromamba install instructions), you can create a new environment using:
micromamba create -n crystall python=3.12
micromamba activate crystall
Then install crystALL (regular)
pip install crystall-searchor if you work with the source code directly, you can clone the GitHub repository and install it in editable mode:
pip install -e .Note: To run certain functions within the crystALL framework, you may need to connect to (a) access-restricted MPDD endpoint (Material-Property-Descriptor Database) or your local MongoDB instance with the data loaded and (b) OPTIMADE-compliant endpoints, which are all open and include MPDD data with minor delay, for remote crystal structure data.
The dgl library is infrequently updated as of 2026 but still widely used for deep learning on graphs and required by ALIGNN models within the crystALL framework, which are currently tightly integrated (until v0.6.0, which is fully modularized, becomes live). If you are having trouble installing torch / dgl / mpdd-alignn on your system, the recommended approach is to:
-
Consider utilizing the provided pre-configured GitHub Codespace, which includes all
crystALLdependencies and a local MongoDB instance you can populate with MPDD or any other database of your choice, to avoid installation issues on your local system -
If you are using Linux, the recommended approach is to first install the required system dependencies via wheels directly from
pytorch.organd then fromdgl.aiwith the following commands:pip install --extra-index-url https://download.pytorch.org/whl/cpu "torch==2.7.0+cpu" "torchvision==0.22.0+cpu" "torchdata==0.9.0" pip install --no-deps 'dgl==2.5.0' -f https://data.dgl.ai/wheels/torch-2.6/repo.html
-
If you are using Windows or macOS, the versions between
v0.3.xandv0.5.0should install without major issues using standardpipcommands, possibly with manualpip install "dgl==2.2.0" -f https://data.dgl.ai/wheels/torch-2.3/repo.htmlfirst; however, versionsv0.5.xhad to keep up with the latest releases oftorch. -
If you need to install the latest version on Windows or macOS, consider doing so in environments like Windows Subsystem for Linux
WSL, Linux VM, orDockerto avoid compatibility issues.
If you would like to try crystALL's functionalities in a reproducible, hands-on environment, we provide a pre-configured GitHub Codespace with all required dependencies and local-access to MongoDB database for running our tutorials:

- Click the button above to launch a ready-to-use, cloud-based workspace.
- Once your Codespace loads, navigate to the
tutorialdirectory to find curated Jupyter notebooks illustrating the main features and scientific workflows. - Open any notebook and execute cells interactively to explore candidate structure generation, ML-powered predictions, and data-driven analysis.
The tutorials currently consist of 3 Jupyter notebooks covering:
- 01. QuickStart.ipynb - Core API, BaSO4 walk-through, clustering and visualization
- 02. HydridesExample.ipynb - Real-world discovery in complex metal hydrides.
- 03. RefiningMPDDSearch.ipynb - Advanced queries (dimensionality filtering, crystal system constraints, compositional limits).
The upcoming v0.6.0 release introduces a fully modular architecture driven by the MLIPAdapter registry, enabling automated structure relaxations and multi-property predictions across diverse machine-learned interatomic potentials. Each potential manages its own execution micro-environment with support for CPU and CUDA acceleration, full cell/volume optimization, and optional piping into downstream static property models. Below is a summary of the machine-learned interatomic potentials integrated into crystALL v0.6.0 via the MLIPAdapter registry:
| Model Family | Checkpoint Pinned | Hardware | Distribution | Citation |
|---|---|---|---|---|
| DeePMD | DPA-4.0.1-Pro-MPtrj | CPU, CUDA | From source | Wang et al. (2018) |
| CHGNet | 0.3.0 | CPU, CUDA | PyPI | Deng et al. (2023) |
| TensorNet | PES-MatPES-PBE-2025.2 | CPU, CUDA | PyPI | Simeon & de Fabritiis (2023) |
| ORB | v3-omat:float32-high | CPU, CUDA | PyPI | Rhodes et al. (2025) |
| ORB-D3 | v3-omat:float32-high:PBE:BJ | CPU, CUDA | PyPI | Rhodes et al. (2025) |
| Allegro | OAM-L:0.1 | CUDA | From source | Musaelian et al. (2023) |
| AlphaNet | alex-0410:oma | CPU, CUDA | From source | Yin et al. (2025) |
| Eqnorm | eqnorm-mptrj | CPU, CUDA | From source | Chen et al. (2025) |
| eSEN | eSEN-30M-OAM | CPU, CUDA | From source | Fu et al. (2025) |
| MACE | mh-1:omat-pbe | CPU, CUDA | PyPI | Batatia et al. (2022) |
| MatterSim | v1.0.0-5M | CPU, CUDA | PyPI | Yang et al. (2024) |
| NequIP | OAM-XL:0.1 | CUDA | PyPI | Batzner et al. (2022) |
| Nequix | oam-1 | CPU, CUDA | From source | Koker et al. (2025) |
| PolarMACE | polar-1-l | CPU, CUDA | From source | Batatia et al. (2026) |
| QET | PES-MatPES-PBE-2025.2 | CPU, CUDA | From source | Ko et al. (2025) |
| SevenNet | omni-i12:omat24 | CPU, CUDA | PyPI | Park et al. (2024) |
| UPET | oam-xl | CPU, CUDA | PyPI | Bigi et al. (2026) |
| EquiformerV3 | omat24-mptrj-salex-gradient | CPU, CUDA | From source | Liao et al. (2026) |
| MatRIS | 10m-oam:efsm | CPU, CUDA | From source | Zhou et al. (2025) |
| UMA | s-1p2:omat | CPU, CUDA | PyPI | Wood et al. (2025) |
| GRACE | 2L-SMAX-OMAT-large | CPU, CUDA | PyPI | Bochkarev et al. (2024) |
| TACE | OAM-L | CPU, CUDA | From source | Xu et al. (2025) |
| HIENet | 0 | CPU, CUDA | From source | Yan et al. (2025) |
| OSAKA | osaka26:small | CPU, CUDA | PyPI | Kuroda et al. (2026) |
| Total | 24 Models |
We welcome contributions from the wider scientific community and appreciate bug reports/fixes, documentation improvements, new adapters/models, and larger feature proposals.
For coding contributions, fork the repository, create a topic branch, and open a pull request (PR) against main. Please follow the project's linters/formatters (Black/flake8/isort), add type hints and clear docstrings for public APIs, include pytest tests (prefer existing fixtures), and avoid adding large dependencies without prior discussion.
If reporting a bug or requesting a feature, please open an issue with a minimal reproduction (code snippet or dataset), environment details, and the expected vs actual behaviour; users may also raise questions and contribute to discussions via issues.
Adding property-prediction models via calculator adapters:
Extending crystALL with new property predictors is welcome. We integrate models as calculator adapters so the rest of the codebase can call different predictor objectives via a uniform interface (see alignn.py and chgnet.py). Key requirements for adapters:
- Adapter location: Create a module under
crystall/calculators/(one file or a small package). - I/O contract: accept pymatgen
Structure; document outputs in the module docstring. - API methods: adapters must expose two public methods:
run_on_structure— run the model on a single structure and return a value or structured dict.run_on_directory— run the adapter over many structures and support both serial and parallel execution.
- Tests & examples: include unit tests with tiny fixtures.
- Lightweight runtime: Avoid forcing heavy runtime dependencies for users who don't need them. If an adapter requires optional packages, guard imports with try/except and show a clear error message describing how to install the extras.
crystALL is an open-source Python package developed by the Phases Research Lab at The Pennsylvania State University, under the supervision of Dr. Zi-Kui Liu, and distributed under the MIT License.
If you use crystALL in scientific publications, please cite:
- Adam M. Krajewski, Ricardo Amaral, Jonathan W. Siegel, Luke A. Myers, Rushi Gong, Shun-Li Shang, Zi-Kui Liu, "Data-driven candidate structure ensemble generation beyond the convex hull with crystALL approach" (October 2026)